Paracetamol
Tested against md-tools 0.6.1. A single small molecule in water: 20 atoms, one rotatable
amide, one ring. The system to use when the question is about a LIGAND rather than about a protein.
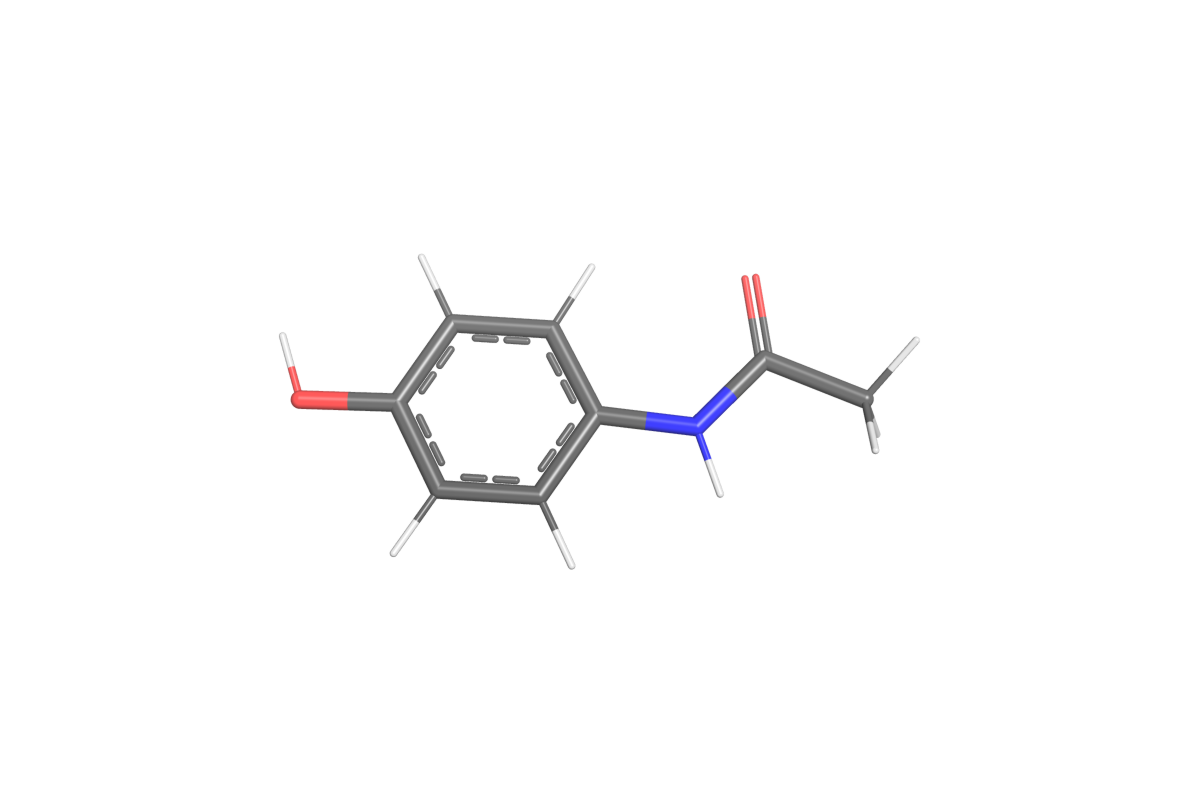
Paracetamol (acetaminophen), embedded and minimised from its SMILES.
Why this system
A ligand needs parameters before it can be simulated, and paracetamol is small enough that parameterising it takes about a minute — so it is the system where the parameter package, its identity and its reuse can be shown end to end without waiting.
Build it
Unlike a protein, a small molecule has no library of residue templates, so the first step produces a parameter package of its own:
md-openmm build-top --parameterize -i paracetamol.sdf --config para.config \
    -op build/parameter/TYL.pdb -os build/parameter/TYL.xml \
    -log build/parameterize.log --resname TYL
-i reads a .sdf, a .mol2 or a .smi. A .smi states the chemistry and no coordinates, so a
conformer is embedded; a structure without bond orders, such as a .pdb, is refused, because
parameterisation needs the molecular graph.
Then the box, reusing that package rather than parameterising again. build.config:
solute:
  kind: ligand
  parameters: ./build/parameter     # a PATH, relative to this file
solvent:
  model: TIP3P
  padding_nm: 1.0
md-openmm build-top -i build/parameter/TYL.sdf --config build.config \
    -os build/built.xml -op build/built.pdb -log build/built.log
That writes the three files every method on this page starts from: the serialised System, the
structure it matches, and the build record. built.log says reused (stated reference) rather
than created — the minute of charge generation happened once, above, and no later build of this
molecule repeats it.
parameters takes a path, as here, so nothing has to be registered and no catalog has to exist.
Give <compound-id>/<parameter-id> instead to take a package from the catalog by name, search
(the default) to find one whose recorded criteria match this build, or generate to parameterise
regardless of what the catalog holds.
Implicit solvent replaces the solvent block and produces no box at all:
The package is identified by its chemical state and its parameters, not by the conformer it was made from, so the same molecule from a different pose resolves to the same package.
Where to go next
| method | what it is for | |
|---|---|---|
| cMD | parameterise once, then a plain simulation from that package | published |
| REST2 | solute tempering of the molecule itself | published |
| AIS | annealed importance sampling, with torsion reweighting | published |
| Umbrella sampling | a potential of mean force along a chosen torsion | page planned |
Umbrella sampling is listed so the shape of the set is visible; it is not written yet, and it is not linked to a page that does not exist.